In-vitro Diagnostics Regulation correctly applied: These requirements result for your CE-compliant product
You can affix the CE mark and sign the declaration of conformity without worries.
IVDR - Regulation on in vitro diagnostics (EU) 2017/746
Willy Lebherz
Director & founder of easyCE GmbH
Declaration of conformity done!
Receive your CE declaration of conformity ready for signature.
Risk assessment completed!
CE mandatory risk analysis conducted.
Relevant directives & standards applied!
Your product evaluated against relevant directives and standards.
Technical documentation created!
Receive a pragmatic, user-friendly operating manual.
CE mark attached!
Simply affix the CE mark and you're done.
CE marking routines differ depending on your role
CE AS PRODUCER
Before 2022, three directives regulated medical devices in the EEA: the Active Implantable Medical Devices Directive (AIMDD - 90/383/EEC), the (General) Medical Devices Directive (MDD - 93/42/EEC) and the In Vitro Diagnostics Directive (IVDMDD - 98/79/EC).
The new EU Regulation on in vitro diagnostics (IVDR), Regulation (EU) 2017/746, replaced Directive 98/79/EC. It modernises the legal framework for medical devices and sets stricter requirements. Manufacturers had a transitional period until May 2022 to adjust their documentation and processes.
According to the Regulation (2017/746), in vitro diagnostics (IVDs) are medical devices that examine human body samples outside the body. They provide information on health conditions, genetic aspects, treatment effects and monitoring of therapies. Examples include HIV test kits, blood glucose meters and special sample containers for diagnostic purposes outside the body. General laboratory products are not covered by this category.
IVDR - Regulation on in vitro diagnostics (EU) 2017/746
The experts at easyCE will be happy to help you place your products on the market and assess special cases. We are happy to support you with CE, UKCA and other labelling.
FAQ
Which EU directives applied to medical devices before 2022?
Medical devices were regulated by three directives in the EEA (European Economic Area) until May 2022.
- Directive on active implantable medical devices, AIMDD (90/383/EEC)
- (General) Medical Device Directive, MDD (93/42/EEC)
- Directive on in vitro diagnostics, IVDMDD (98/79/EC)
What is the new EU regulation on in vitro diagnostics and its transitional period?
Two new regulations now replace the above-mentioned directives on medical devices in order to:
- Create a modernized and more robust EU legal framework,
- Remain at the cutting edge of science and technology, and
- Provide better protection of public health and patient safety.
The new Regulation (EU) 2017/746 (hereinafter referred to as IVDR) on in vitro diagnostics replaces Directive 98/79/EC. Manufacturers have a transitional period of five years until May 2022 to update their technical documentation and processes to meet the new, more stringent requirements.
What are in vitro medical devices (IVDs)?
The new regulation (2017/746) defines an in vitro diagnostic agent (IVD) as: "medical device used as a reagent, reagent product, calibrator, control material, kit, instrument, apparatus, software or system - individually or in conjunction with each other - by the manufacturer for in vitro testing of samples originating from the human body, including blood and tissue donations, is intended and is intended solely or primarily to provide information on one or more of the following
- Physiological or pathological processes or states,
- Congenital physical or mental impairments,
- Predisposition to a particular health condition or disease
- Safety and tolerability of potential recipients,
- Expected effect of treatment or likely reactions to it,
- Determination or monitoring of therapeutic measures
Examples of IVDs are:
- HIV test kits
- Antigen tests
- Immunoassay analyzers
- Blood gas analyzers
- Calibrators and control materials used to check the performance of the analyzers
- Sample tubes and blood collection tubes
- Blood glucose meters and strips
Products for general laboratory use are not considered IVD, but products specifically designed for in vitro diagnostic testing are considered in vitro diagnostics.
What is the category of medical in vitro diagnostics (IVD) under the old directive?
In the IVD Directive (98/79/EC), IVDs are hierarchically divided into the following four categories according to perceived risk:
- General/other IVDs: Products other than those covered by Annex II to the Directive and products for own use
- IVDs for self-testing: Products that may be used by laypersons in the home environment, except for the self-testing equipment referred to in Annex II
- IVDs according to Annex II List B: Reagents and products for rubella, toxoplasmosis and phenylketonuria as well as devices for self-monitoring of blood glucose
- IVDs according to Annex II List A: Reagents and products for human immunodeficiency virus I and II and hepatitis B, C and D
IVD Directive 98/79/EC (IVDD) requires the inclusion of BS/CA only for a few high-risk IVD listed in Annex VII. easyCE can help you classify your product.
What is the new classification for IVDs under IVDR?
In a slight deviation from the IVDD, the IVDR introduces a risk-based classification of classes A to D, where D represents the highest and A the lowest risk category.
This new classification means that for most IVDs, a notified body or a competent authority must be included in the CE conformity assessment process (as opposed to the manufacturer's self-declaration under the IVDD).
What applies to IVD class D products according to IVDR?
The following risk conditions generally apply to Class D products:
- general life-threatening conditions
- transmissible agents in blood and biological materials intended for transplantation or reprocessing into the body (transmissible agents may present a high risk)
- Blood grouping or tissue typing, if they are markers of the following systems: ABO, Rhesus, Kell, Kidd and Duffy
What applies to IVD class C products according to IVDR?
Class C includes the following risk conditions:
- high-risk IVD products that pose a lower risk to the general public
- if the failure of a diagnosis could be life-threatening (including tests for infectious diseases and cancer)
- accompanying diagnostics and genetic screening
- own application in general with certain exceptions.
What applies to IVD class B products according to IVDR?
Class B covers all IVD products that are not specifically covered in other classification rules. These usually include the following products:
- Products with lower risk to the patient and the population than IVD Class D and C products
- self-medication for pregnancy and fertility tests as well as for the determination of cholesterol levels and for the detection of glucose, erythrocytes, leukocytes and bacteria in the urine
- Controls without a quantitative or qualitative value.
What applies to IVD class A products according to IVDR?
Class A comprises laboratory equipment, instruments and sample vessels in the broadest sense. Only Class A devices are approved for self-certification for conformity assessment with the CE marking.
What does the new classification of IVD products mean?
Seven rules describe risk-based IVDR classes that take into account the intended purpose of the products and their inherent risks. In classification, the emphasis is on demonstrating the performance of the product.
Although most IVD medical devices must be certified under the new classification rules by notified bodies (BS or NB) (an estimated 80% of manufacturers are classified into higher risk classes according to IVDR)this is essential for improving patient safety. Nevertheless, the manufacturers of these products will be heavily affected by the complex requirements and the control by the notified bodies.
What are the challenges of the new IVDR?
Important aspects of Regulation (EU) 2017/746 on in vitro diagnostics (IVDR) that are challenging for manufacturers:
- classification rules
- technical documentation
- general safety and performance requirements
- unique product marking (UDI)
- EUDAMED
- performance evaluation
- monitoring after placing on the market
How can easyCE help me? The easyCE expert network supports you in the creation or conversion of necessary documents on all points listed above, for example. easyCE also advises you on the necessary information on the product, the packaging and the package insert.
What innovations does the IVDR bring with regard to technical documentation?
The requirements and the expected quality of the technical documentation have increased. The documents are subject to a closer examination by the competent authority (NL). The clinical data must be solid enough to properly substantiate all claims. Manufacturers must improve the scientific quality and comprehensibility of their technical documentation.
The technical documentation for IVD shall include information on the design, intended use, risk assessment and path to conformity in accordance with general safety and performance requirements and the quality management system (QMS). The post-market performance assessment and monitoring report shall be kept and updated in the technical documentation in addition to the design history record and.
How do I archive the technical documentation?
Technical documentation should be kept secure and protected from unauthorized access and alteration. Under IVDR legislation, manufacturers are required to keep the technical documentation for CAs/NBs for a period of at least 10 years after the last product is placed on the market.
What innovations does the IVDR bring regarding the general safety and performance principles (SPR)?
The extended version of the Basic Requirements is now referred to as the General Safety and Performance Principles or Requirements (SPR). The SPR/GSPR under the IVDR cover many aspects to be considered and are listed in Annex 1 to the Directive. easyCE can support you in the preparation of the necessary documentation.
What is the UDI system?
The introduction of an UDI system aims to improve the traceability of all medical devices on the market by placing a unique code on the label of the product. UDI is primarily used for reporting serious incidents and identifying counterfeit medical devices.
According to the IVDR regulation, every manufacturer is obliged to introduce a UDI system. The basic UDI-DI must be included in the technical documentation, the declaration of conformity and the certificate of the notified body. Basic UDI-DI and UDI-DI are used for registration in EUDAMED.
What is EUDAMED?
As part of the IVDR/MDR, EUDAMED serves as a central database for the registration of economic operators and products, but also for vigilance reports and performance studies. EUDAMED will improve the transparency and coordination of information on medical devices available on the EU market to meet vigilance reporting and analysis requirements, including serious incidents On-site safety corrective actions and trend reports.
What innovations does the IVDR bring with regard to performance evaluation?
A major challenge of the IVDR is the updated and specified requirements for performance evaluation. It defines a continuous process to provide sufficient clinical evidence to demonstrate that the intended clinical benefit is achieved and the product is safe.
What innovations does the IVDR bring with regard to post-market monitoring?
PMS and in this context Post-Market Performance Follow-Up (PMPF) play an important role within the IVDR. Post-market data must be used to update essential documents, such as risk management and performance evaluation. Manufacturers are required to prepare the relevant documents in accordance with the IVDR, for example the PMS Plan and Report/ Periodic Safety Update Report (PSUR).
How is the CE marking procedure described under the IVDR?
The CE marking procedure for in vitro diagnostics covers two aspects: general compliance and the actual conformity assessment procedure. easyCE accompanies you through the entire procedure outlined below:
- Appointment of a European representative
- Classification of the device and identification of all EU directives/regulations applicable to your product (such as IVDR, WEEE, EMC, etc.)
- Selection and implementation of the product-specific conformity assessment procedure
- Verification of the equipment against general safety and performance requirements (Annex I)
- Identification of applicable harmonised standards
- Preparation of a technical dossier (Annex II and III) and implementation of a quality management system, preferably based on ISO 13485
- Selection of a notified body (except for Class A)
- Audit of the QMS and technical documentation/design dossier by a notified body
- CE marking and ISO 13485 certificates issued by the notified body
- Preparation of a legally binding declaration of conformity (DoC) by the manufacturer
- Affix the CE marking
- Registration of IVD with the competent European authority (Ministry of Health), in the country of the authorised representative
How does the conformity assessment procedure for IVD differ from other medical devices?
The CE conformity assessment procedure for IVD is similar to that for medical devices, but there are some significant differences due to the different risks associated with the different products. This results in different conformity paths for different risk-based classes.
How do I get an ISO 13485 certification?
For products other than "A", the QMS and technical documentation must be audited by a European notified body. The CE marking certificate of the device and the ISO 13485 certificate of the production site are issued after successful completion of the 3-party audit. The ISO 13485 certification must be renewed every year, CE marking certificates are usually valid for 3 years.
How are the harmonized IVDR standards developing?
On 15 May 2020, the European Commission published its Implementing Decision M/565 on the development of harmonized standards under the Medical Devices Regulation (MDR) and the In Vitro Diagnostics Regulation (IVDR). Implementing Decision M/565 reflects significant changes in the development of harmonized standards for IVD products:
- The changes are technology-neutral and performance-related.
- Most standards refer to processes and not products and are harmonized.
- The number of harmonized standards is decreasing, there are only 41 for IVDs.
- Standards should address several specific aspects, e.g.:
- EN ISO 10993-7:2008: The method for calculating the maximum residue limits should also consider the body weight of well below 70 kg
- EN ISO 10993-17:2009: Biological assessment of medical devices - Part 17: Determination of permissible limit values for excreted substances Body weights below 3.5 kg (newborns) must be taken into account
- ISO 23908: Protection against injuries caused by sharp/pointed instruments - Requirements and test methods - Features for protection against sharp/pointed instruments for subcutaneous needles, insertion tools for catheters and needles for blood collection: MDR/IVDR Annex I, Sections 11.1 (Reduction of the risk of infection for patients and users) and 22.2 (products for lay use) must be taken into account in the harmonized version of ISO 23908.
What are the tasks of the responsible person/expert?
Article 15 of the IVDR clearly states that manufacturers of medical devices are obliged to have within their organisation or at their disposal at least one person who has the necessary expertise in the field of medical devices. The responsible person must ensure that the technical documentation is prepared and maintained.
What are the tasks of the authorized representative?
In accordance with Article 11 "the manufacturer of a product not established in a Member State shall designate a single authorized representative". The authorized representative shall provide a copy of the technical documentation, thereby ensuring effective communication with the notified bodies in relation to the manufacturer's obligations.
What are Notified Bodies for?
Notified bodies (BS) or Notified Bodies (NB) are responsible for ensuring the safety and conformity of all in vitro medical devices placed on the EU market.
Hundreds of companies trust easyCE



With easyCE, product compliance is really 'easy'
Approach to CE marking
Type and manner of support
Scope of support
Contact during project processing
 Find the easiest way to obtain the CE mark
Abbreviated process for CE derived directly from CE
All-inclusive or modular – whatever you prefer
Personal contact with experts who take your wishes into account
other CE service providers
 Often lacking in sound judgement and unnecessarily complicated
Mostly flexible but often bureaucratic
Mostly flexible in terms of support scope
Mostly personal contact
CE Software
Follows standardised process; special circumstances not taken into account
Double burden for you: software management + no support during processing
Only according to software scope and purchased modules
No contact
You want support that lightens your workload and doesn't create more work.
This is what successful projects look like
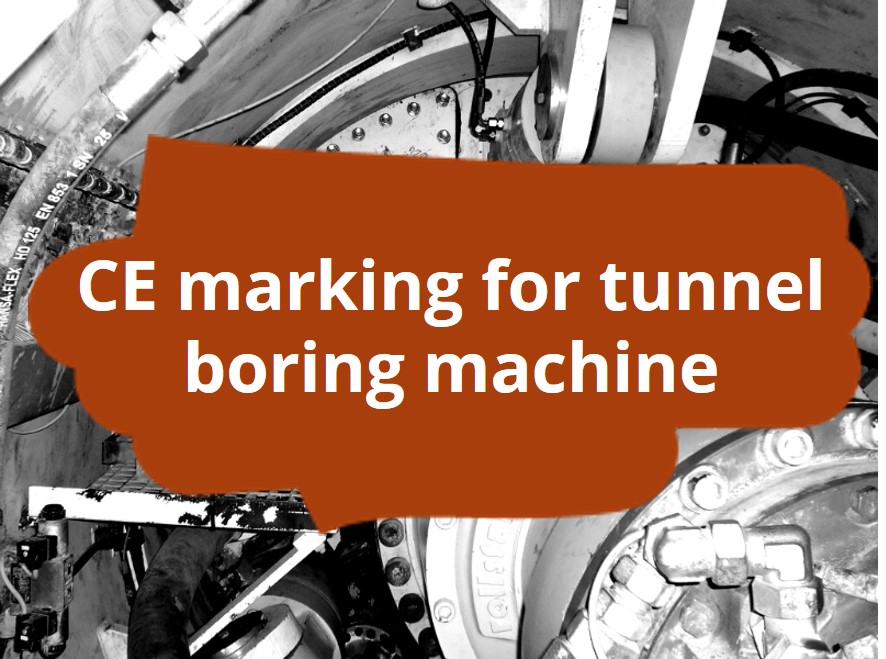
CE marking and conformity assessment of all life phases of a copper mining tunnel boring machine in Australia
easyCE supports the entire conformity assessment process including risk assessment up to the declaration of conformity ready for signature for all phases of the life cycle of a tunnelling machine for copper mining in Australia. The completely new concept of using multi-arm cutting heads required special attention during the process. In addition to the conventional regulations, special safety and technical requirements that apply in Australia were also taken into account.
RL & VO: MRL - 2006/30/EU, EMC - 2014/30/EU, Low voltage - 2014/35/EU, PED - 2014/68/EU, ATEX - 2014/34/EU
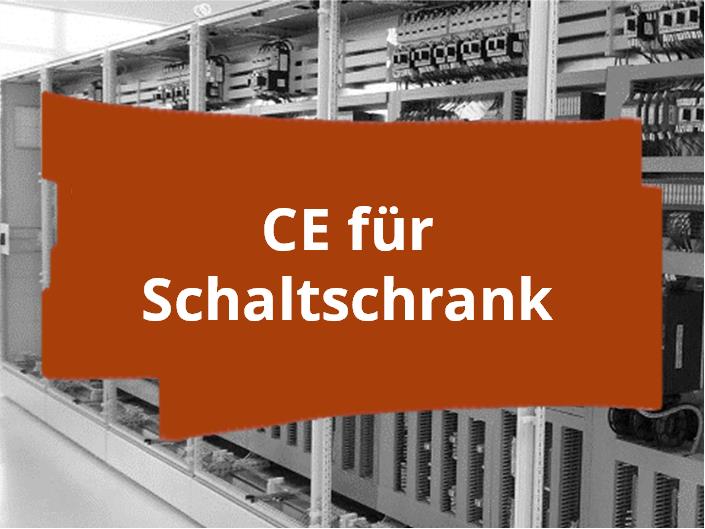
CE labelling of switch cabinets
easyCE supports the entire conformity assessment process, including risk assessment, right through to the declaration of conformity for control cabinets ready for signature. In particular, the requirements from the Machinery, EMC and Low Voltage Directives with regard to DIN EN 60204-1: 2019-06 and DIN EN IEC 61439-1: 2021-10 had to be observed. A checklist was also drawn up for the selection of components that must fulfil special requirements (supplier selection).
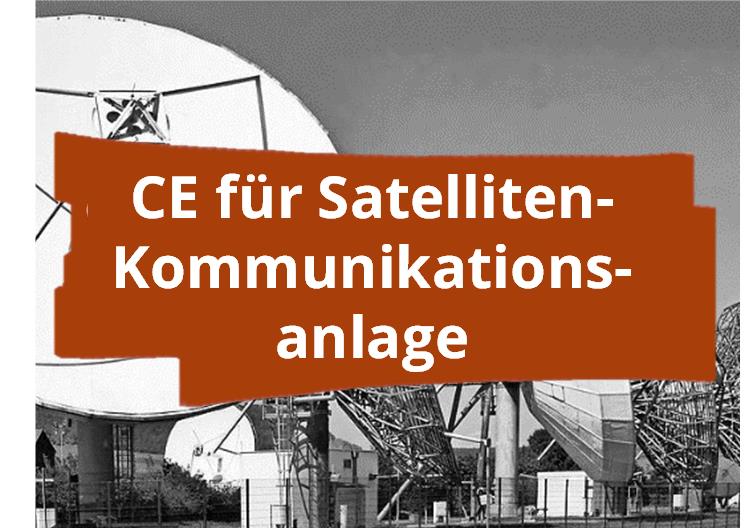
CE marking and declaration of conformity of a mobile satellite communication system
easyCE supports the entire conformity assessment procedure, including risk assessment in accordance with the EMC Directive, right up to the ready-to-sign declaration of conformity for a satellite communication system. The system ensures the redundant forwarding of TCP/IP data in order to send or receive it via two or more satellite antennas. The system is used wherever a mobile communication system needs to be set up quickly over long distances.
RL & VO: RED - 2014/53/EU, Low Voltage - 2014/35/EU, EMC - 2014/30/EU
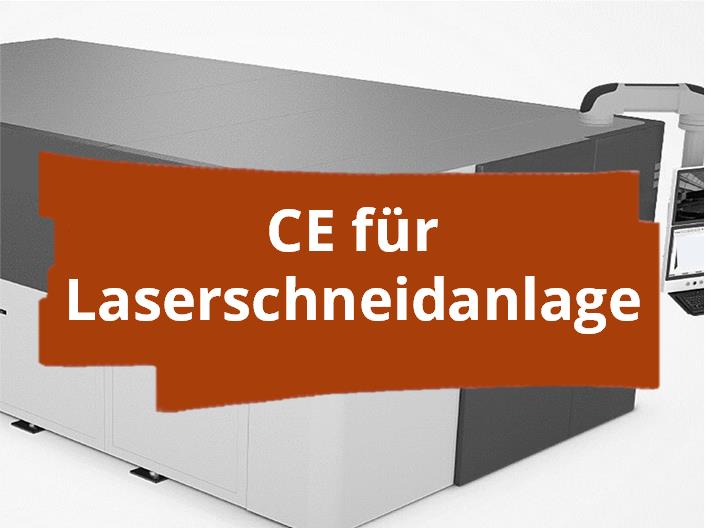
CE marking and declaration of conformity for a 2D laser cutting system
easyCE supports the entire conformity assessment procedure for a 2D laser cutting system with automatic pallet changer. The entire safety concept, including the 2-beam light barrier, was coordinated for the laser cutting system, the performance level was determined, a product analysis was created and a risk assessment was carried out in accordance with EN ISO 12100. Special requirements result for the mechanical components of the traverse for the laser cutting head, but especially for the kinematics of the pallet changer and laser cutting head. The cutting process (laser class 4) inside the cabin is safely shielded from the operator by laser protection screens, additional safety devices and the complete enclosure: laser class 1 outside the machine.
Takes into account among others: 2006/42/EC, 2014/30/EU, DIN EN 60825-1 etc.
See more of our project references and success stories
Get CE-marking done in 3 simple steps
Step 1
Request expert opinion
Ask for our expert opinion free of charge and find out what is necessary for the CE marking of your product. Be prepared to describe your product in broad terms.
Step 2
Receive our offer
If your product is subject to CE marking, we offer you our support with the CE marking and all accompanying measures. We will be happy to explain necessary steps on the phone.
Step 3
We get it done
When you place an order, you will be assigned your own personal CE expert who will carry out your CE marking and accompany you throughout the entire process. Your expert will also be at your side after the CE marking has been successfully completed.
Get CE-marking done, worry-free
Trust the experts of product safety and conformity
Willy Lebherz - Founder and Managing Director of easyCE
- Expert in product safety and conformity since 1995
- Recipient of the "Medal of the Order of Merit of the Federal Republic of Germany", awarded in 1983 by the then Federal President Carl Carstens
- Master of measurement and control technology
- Captain (ret.), Project Officer for Technical Logistics in the Army Material Office and Chief of the Telecommunications Repair Company
easyCE is a digital, dynamic engineering office with a focus on product safety and product conformity - especially CE-marking. We support manufacturers, operators and dealers in designing products safely and offering them on the market in compliance. We are a "full-service provider" and can take over the entire conformity assessment process for you if you wish. To do this, we support you in carrying out risk assessments, researching standards, product analyses, developing suitable safety precautions, preparing user-friendly technical documentation, coordinating tests and all other accompanying measures. easyCE was founded in southern Germany, but is now active globally.
We know what you need to do to get CE-marking done, worry-free.