The technical documentation of the Medical Devices Regulation
You can affix the CE mark and sign the declaration of conformity without worries.
Willy Lebherz
Director & founder of easyCE GmbH
Declaration of conformity done!
Receive your CE declaration of conformity ready for signature.
Risk assessment completed!
CE mandatory risk analysis conducted.
Relevant directives & standards applied!
Your product evaluated against relevant directives and standards.
Technical documentation created!
Receive a pragmatic, user-friendly operating manual.
CE mark attached!
Simply affix the CE mark and you're done.
CE marking routines differ depending on your role
CE AS PRODUCER
The MDR 2017/745 requires new standards for the technical documentation of medical devices and requires a conversion from traditional to MDR-compliant approaches. Key points are the Design History File (DHF), which documents development processes, and the technical dossier, which includes clinical evaluation and post-market studies. It includes information on product development, verification, clinical performance and market monitoring to continuously meet regulatory requirements. easyCE offers help with the creation of MDR-compliant documentation and individual consulting for required documents depending on the product type.
Find out here what the requirements of the EU Directive 2011/65/EU are and how this can influence the CE marking of your product.
The experts at easyCE will be happy to assist you in placing your product on the market in the EEA and in assessing special cases. We support you with CE, UKCA and other markings.
FAQ
How does the technical documentation change with the new MDR (2017/745)?
The latest EU Regulation 2017/745 for medical devices (hereinafter referred to as MDR) provides a detailed framework for correct technical documentation. This article will mainly refer to the MDR as it provides
a comprehensive understanding of the technical documentation required in the near future. Manufacturers must move from the traditional approach of technical documentation to the new concept of the new regulation. The easyCE expert network is happy to support you in the conversion of your technical documentation and in achieving CE conformity of a new product.
What is Design History File (DHF) according to MDR?
DIN EN ISO 13485:2021-12 defines the DHF by the necessity to create "design and development files" as follows:
"The organization must have a design and development file for each type of medical device or each medical product family. This file shall contain or refer to records prepared to demonstrate conformity with the requirements for interpretation and development, as well as records of changes in interpretation and development."
The DHF is now the key file that documents design checks. the DHF is a collection of documents from the design and development process. Thus, the DHF verifies the product development process and proves that the product is effective and safe. The DHF is a "living file" that is updated throughout the life cycle of the product. Changes made during or after production and the associated design checks must be stored in the DHF. The risk assessment must be adjusted with each change.
What is a medical device technical dossier?
The technical dossier (also: technical documentation/file) documents the manufacturing process and proves that the product is safe and effective and can therefore be sold in the EU. In order to create a technical dossier correctly, the verification and design validation processes documented in the DHF are required.
In addition to the DHF, a technical dossier must also contain a report on the clinical evaluation. It deals with the clinical trial of the product, the risks and any clinical follow-up studies that may be required after placing it on the market.
What does the technical documentation according to the MDR for medical devices contain?
The technical documentation is the entirety of the documents describing your product, which are to be understood only as indications. Further documents may be required to comply with the regulation.
The technical information shall include at least information on the design, development, verification and validation of clinical performance and the approval status of the product in the target markets. In addition, the MDR requires post-market monitoring of the product (PMS) to ensure a closed loop. This ensures the early detection of hazards, so that the "General Safety and Performance Requirements" (GSPR - according to ANNEX 1 of the MDR) are continuously met. An appropriate risk management system ensures that the benefits outweigh the risks. You will find a list of the minimum content of the technical documentation in accordance with Annexes II (a) and III (b) to the MDR below.
- Product description and specification, including variants and accessories
- Product description and specification
- Reference to previous and similar generations of the product
- Manufacturer information on labelling, handling conditions, sales states and instructions for use
- Information on design and production
- General safety and performance requirements
- Benefit-risk analysis and risk management
- Product review and validation
- Preclinical and clinical data
- Additional information required in special cases
- PMS plan and reports (post market surveillance = post-market surveillance)
- PSUR (Periodic Safety Update Report = regularly updated safety report)
How do I create MDR-compliant technical documentation with DHF?
Appendix II of the MDR contains instructions for creating and maintaining the technical documentation. In practice, in accordance with Annex II, the DHF includes the following documents:
- Input requirements and specifications, purpose, mode of operation, variants
- BASIS-UDI-DI and EUDAMED registration
- Design drawings
- Results of design review according to company-specific development protocol
- System for traceability of the design process
- Component testing information (test plan and data)
- Information on biocompatibility of materials (test plan and data)
- Electrical/electromagnetic safety tests (if applicable)
- Further verification and validation results (identification, methods, data, people, etc.) for the first finished devices
- Documentation of studies to prove conformity (e.g., laboratory tests, animal tests, literature searches for similar products)
- Software verification and validation (if applicable)
- Risk management plan
- Sterilization information (if applicable)
- Information on the manufacturing process (suppliers, sub-contractors, work instructions, auxiliary materials)
- Product photos and sketches
- User information such as package inserts and packaging with information on shelf life and storage conditions
- List of harmonized standards applied
Do not hesitate to contact easyCE's expert network. We are always available to assist you in the preparation and compilation of these documents. In addition, we are happy to offer you individual advice to determine which documents are required specifically for your product.
How do UDI-DI and basic UDI-DI differ?
For EU-wide product identification and traceability, the MDR requires a unique product identification of medical devices. In practice, this is implemented by the UDI-DI (Unique Device Identifier - Device Identifier) and UDI-PR
(Unique Device Identifier - Production Identifier). While the UDI-DI is unique, a basic UDI-DI can refer to several products. However, all products with the same basis UDI-DI have the same core characteristics, such as purpose, risk class, essential design and manufacturing characteristics. A basic UDI DI always refers to at least one UDI DI, while several UDI DI can refer to the same basic UDI DI.
What are SRN and EUDAMED registration?
Rie traceability of UDI-DIs is done by EUDAMED. Create an account with EUDAMED online and you will receive an SRN (Single Registration Number) after verification. In EUDAMED, store the UDI/product information of all products that you place on the EU market. However, EUDAMED has initial difficulties, which is why you have to pass on additional information to state authorities for the time being.
What does STED mean for medical devices?
The Global Harmonization Task Force (GHTF) created a standardized and summarized form of documentation, the "Summary Technical Documentation (STED)". This is sufficiently detailed to enable a notified body (BS or NB) to fulfil its obligations. The International Medical Devices Regulators Forum (IMDRF) then initiated a guideline providing for an internationally harmonized table of contents to be used for electronic submissions in the future.
What does dynamic documentation mean for medical devices?
The technical documentation is intended to be a living document that must be created during the design and development process of a device and maintained and regularly updated throughout its life cycle.
What is meant by verification and validation of medical devices?
Design reviews and continuously documented development processes verify the acceptance criteria of your defined milestones. You must keep the documentation you create during the design and development reviews securely in either a Design History File (DHF) or a Device Master Record (DMR).
What is PMS and PMCF for medical devices?
The PMS (Post Market Surveillance) consists of the data recorded after placing on the market. These may include competitor vigilance data, post-market clinical follow-up (PMCF), user feedback and complaints.
These new inputs can provide a further cycle in the design and development process. Documented checks of the constructive changes and the associated corrective and preventive measures are essential evidence for the continuous fulfilment of the GSPR (General Safety and Performance Requirements).
What needs to be considered in the technical documentation of medical devices?
The technical documentation should be structured and presented in such a way as to facilitate its assessment by the notified body (BS or NB). This means that, when compiling technical documentation, it is necessary to maintain an appropriate level of detail while avoiding the inclusion of superfluous details that are not strictly necessary. The notified body requires at least the following documents:
- QMS (quality management system) according to ISO 13485:2016
- Information on the design and production process
- Design History File (design traceability)
- Device master record (reference for production)
- Continuous monitoring via an electronic database (registration with EUDAMED)
The test results and certificates issued by the notified body are also part of the technical documentation.
How do I implement the traceability of technical documentation for medical devices?
Traceability means that a notified body can track your process chain (all phases of production) using your technical documentation. A clear structure of the entire technical documentation enables proof of compliance with
the GSPR (General Safety and Performance Requirements). Forward and backward traceability is required. Forward traceability refers to the application of the medical device; backward to the origin of the product, its manufacture and any suppliers and sub-providers. Overall, the following aspects must be considered:
- Specification of the user request
- Functional requirement specification
- Risk analysis
- Clinical evaluation
- General requirements
Who is the "Responsible Person"?
Article 15 of the MDR stipulates that medical device manufacturers must have at least one person within their organisation or at their disposal who has demonstrable expertise in the field of medical devices. The responsible person must ensure that the technical documentation is prepared and maintained.
Who is the "Agent"?
Pursuant to Article 11, "the manufacturer of a [medical] product not established in a Member State shall designate a single authorized representative". The authorized representative shall, inter alia, provide a copy of the technical documentation, thereby ensuring effective communication with the notified bodies in relation to the manufacturer's obligations.
What do notified bodies do?
Notified bodies are responsible for ensuring the safety and conformity of all products placed on the EU market. If a Class I or IIa product is at low risk, the manufacturer may self-certify. However, an authorized representative is required to legally represent the product on the market.
What are the archiving periods for the technical documentation of medical devices?
You must keep the technical documentation safe and secure from unauthorized access and changes. EU legislation requires you to keep the technical documentation for the notified body for a period of at least 10 years after the last device has been placed on the market; 15 years for implants.
What is required for the CE marking of a medical device?
In order to ensure the CE conformity of a medical device, the technical documentation must be supplemented by the issuance of a declaration of conformity and the preparation of a user manual before the product is placed on the market in the European Union. Even with these steps easyCE can advise you or actively assist you.
How do I create a Declaration of Conformity (DoC) for medical devices?
The declaration of conformity is a declaration by the importer or manufacturer pursuant to Article 19 of the MDR that the product meets the essential requirements of the applicable directives. In accordance with Annex IV of the MDR, the declaration of conformity must include the following information:
- Labelling of the product including trade name and basic UDI-DI
- Product features such as specifications and risk class
- Name and address of the manufacturer/importer and SRN
- List of applicable EN standards or directives
- Identification number of the notified body
- Person responsible
- Name and address of the authorized representative
- Place, date, signature, name and function of the signer
Supervisory authorities may at any time request the DoC to verify compliance with the regulations.
What must the instructions for use (package leaflet) of a medical device contain?
They must provide instructions for each medical device to be placed on the market in the EU. Detailed information can be found in the MDR, Annex I, Chapter III. The instructions for use must include the following information:
- Purpose
- Reference to Notified Body Clinical Performance Summary
- Correct use
- Precautions and safety precautions for residual risks
- Contraindications
- CE-Marking
- For reusable products, instructions on: cleaning, disinfection, sterilization, maintenance and functional testing
- Transport and storage
- Safe disposal
- If necessary, radiation emissions
Regulation (EU) No 207/2012 contains information on electronic instructions for use for medical devices.
How and when is compliance with EN standards required?
Harmonized standards are EN standards whose reference numbers have been published in the Official Journal of the EU. They justify the presumption of conformity with the essential requirements of the various EU regulations. The declaration of conformity shall contain a list of the harmonized standards applied as part of the technical documentation.
Examples of harmonized standards for medical implants published in the EU Official Journal are:
- EN 556-2:2015 "Sterilization of medical devices"
- EN ISO 11137-1:2015 + A2:2019 Sterilization of healthcare products - Blasting - Part 1: Requirements for the development, validation and control of the application of a sterilization process for medical devices"EN ISO 11137-2:2015 Sterilization of healthcare products - Blasting - Part 2: Determination of sterilization dose"
How do I create a risk management plan for my medical device?
The risk management plan is an important part of the technical documentation. The experts at easyCE are very familiar with the new procedure for risk management in medical devices presented in DIN EN ISO 14971:2022-04 and are happy to advise and support them. Repeating the following steps forms the basic framework:
- Risk analysis
- Risk assessment
- Risk governance
- Assessment of the overall risk
- Review of risk management
- Information management during and after production
Hundreds of companies trust easyCE



With easyCE, product compliance is really 'easy'
Approach to CE marking
Type and manner of support
Scope of support
Contact during project processing
 Find the easiest way to obtain the CE mark
Abbreviated process for CE derived directly from CE
All-inclusive or modular – whatever you prefer
Personal contact with experts who take your wishes into account
other CE service providers
 Often lacking in sound judgement and unnecessarily complicated
Mostly flexible but often bureaucratic
Mostly flexible in terms of support scope
Mostly personal contact
CE Software
Follows standardised process; special circumstances not taken into account
Double burden for you: software management + no support during processing
Only according to software scope and purchased modules
No contact
You want support that lightens your workload and doesn't create more work.
This is what successful projects look like.
CE marking and Declaration of Conformity for silicone rubber molded parts in industrial applications
easyCE successfully supports CE marking up to the ready-to-sign Declaration of Conformity for elastic molded parts made of RTV silicone rubber. The special challenge with these products lies in meeting the fire safety requirements according to EN 13501-1 and ensuring chemical resistance according to EN ISO 10993-18. The various silicone types must undergo thorough testing for fire resistance, toxicity, and long-term stability, with particular attention to compliance with strict limits for volatile organic compounds (VOC) according to EN 16516.
Considering: Construction Products Regulation (EU) Nr 305/2011
Conformity assessment procedure and CE marking for an anesthetic gas scavenging system in medical environments
easyCE successfully supports CE marking up to the ready-to-sign declaration of conformity for a system designed to scavenge anesthetic gases. According to EN ISO 80601-2-13, special safety requirements for anesthesia workstations had to be met. The challenge was validating flow rates for effective gas extraction while ensuring electrical safety according to EN 60601-1 for use in medical environments. Through precise validation measurements, the system's consistent performance was demonstrated.
Considering: 903/85/EEC
Conformity assessment, CE marking and instructions for use for a permeate treatment plant
easyCE supports the entire conformity assessment procedure, including the instructions for use for a system for permeate treatment, i.e. the treatment of water as a precursor to dialysis machines. In terms of design, it was important for the system to maintain and achieve a high degree of purity, which also had to be taken into account during the risk assessment.
Requirements research for dental implants
easyCE supports the research of a wide range of technical requirements for a dental implant. In particular, the technical properties with regard to the angle and inclination of the threads, but also materials such as titanium, were to be found out.
Support with the CE marking of horn eyeglass frames (Class I medical device) and sunglasses
In a challenging project, easyCE ensured the conformity of horn eyeglass frames as accessories to medical devices under the Medical Devices Regulation 2017/745 and of sunglasses under the PPE Regulation 2016/425. We extended the technical documentation to ensure that all required documents such as the risk management plan and the post-market surveillance plan are included.
Our team worked closely with the client to ensure all requirements of the Medical Device Regulation were met, including the general safety and performance requirements set out in Annex I. This included a thorough review of the design and manufacturing requirements to ensure the products met the highest standards.
CE marking for a lubricant according to the Medical Devices Directive
With the help of the Medical Device Directive 93/42/EEC, easyCE also finds the relevant requirements in the case of ambiguous definitions, informs the customer of their obligations and creates the necessary CE documentation around the existing quality management system. A risk management plan, checklists for technical documents, for hazard identification and safety-relevant features, for general safety and performance requirements according to MDR Annex I and a declaration of conformity are created product-specifically in German and English.
CE marking for drug tests
In this specific case of a multi-drug test, the decision on CE marking requirements depends on the user group. While a drug test for law enforcement purposes is exempt, a drug test for medical purposes falls under the new Medical Device Regulations (EU) 2017/745 and In Vitro Diagnostics (EU) 2017/746 as a Class C product. In this project, easyCE adapts the schedule to the customer in a particularly flexible manner, as unexpected problems arise during production. Even after completion of the commissioned project (Technical File, Hazard Identification, Identification of Safety-Related Characteristics, GSPR File, Risk Management Plan, Declaration of Conformity) and handover to the customer, design changes to the product occur. easyCE adapts the conformity documents and advises on necessary changes to the technical documents. Finally, easyCE contacts notified bodies on behalf of the customer and accompanies the customer during registration in EUDAMED to obtain SRN and basic UDI-DI.
Takes into account, among others: Regulation (EU) 2017/745; Regulation (EU) 2017/746
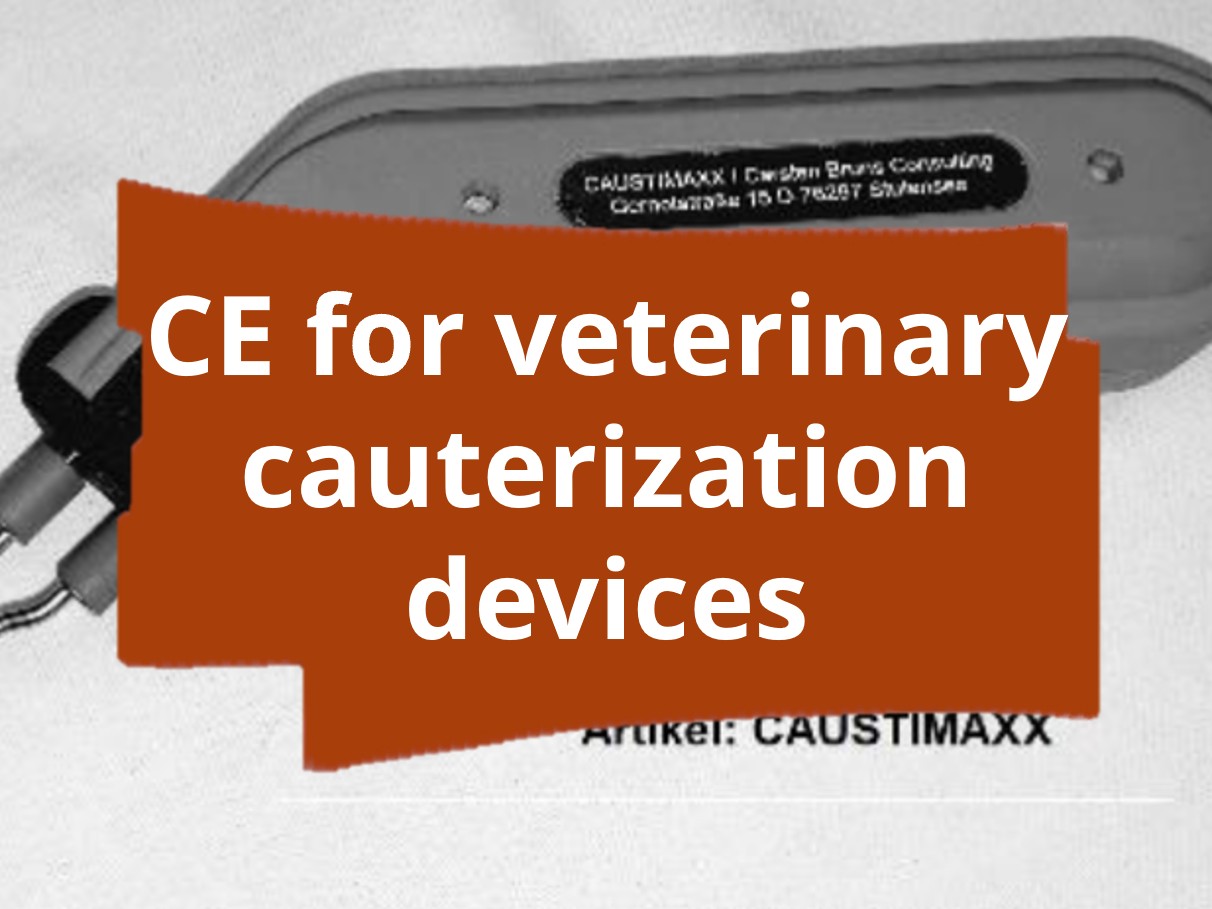
Conformity assessment procedure and CE marking for veterinary cauterization devices
easyCE successfully supports CE marking up to the ready-to-sign Declaration of Conformity for handheld cauterization devices. The implementation of requirements according to EN 60335-1 for electrical safety represents a central aspect. A particular challenge is compliance with electromagnetic compatibility according to the EMC Directive 2014/30/EU, which was solved through targeted measures such as shielding and interference suppression capacitors in accordance with EN 55014-1.
Considering: 2014/35/EU, 2014/30/EU, 2011/65/EU
See more reference projects
Get CE-marking done in 3 simple steps
Step 1
Request expert opinion
Ask for our expert opinion free of charge and find out what is necessary for the CE marking of your product. Be prepared to describe your product in broad terms.
Step 2
Receive our offer
If your product is subject to CE marking, we offer you our support with the CE marking and all accompanying measures. We will be happy to explain necessary steps on the phone.
Step 3
We get it done
When you place an order, you will be assigned your own personal CE expert who will carry out your CE marking and accompany you throughout the entire process. Your expert will also be at your side after the CE marking has been successfully completed.
Get CE-marking done, worry-free
Trust the experts of product safety and conformity
Willy Lebherz - Founder and Managing Director of easyCE
- Expert in product safety and conformity since 1995
- Recipient of the "Medal of the Order of Merit of the Federal Republic of Germany", awarded in 1983 by the then Federal President Carl Carstens
- Master of measurement and control technology
- Captain (ret.), Project Officer for Technical Logistics in the Army Material Office and Chief of the Telecommunications Repair Company
easyCE is a digital, dynamic engineering office with a focus on product safety and product conformity - especially CE-marking. We support manufacturers, operators and dealers in designing products safely and offering them on the market in compliance. We are a "full-service provider" and can take over the entire conformity assessment process for you if you wish. To do this, we support you in carrying out risk assessments, researching standards, product analyses, developing suitable safety precautions, preparing user-friendly technical documentation, coordinating tests and all other accompanying measures. easyCE was founded in southern Germany, but is now active globally.
We know what you need to do to get CE-marking done, worry-free.